A supplement label that says “made in an FDA-registered facility” can sound like a quality seal. It is a narrow statement about a facility’s registration status. It does not tell a customer that FDA inspected the site recently, reviewed a specific batch, endorsed a manufacturer, or approved the product.
A creator or founder who cites registration to build trust can imply more than the record proves. U.S. manufacturers and marketers have legal duties, and FDA conducts inspections and enforcement; those roles do not make facility registration, a quality record, a permitted claim, and product approval interchangeable. A product page that blurs them risks misleading customers and obscuring what the evidence actually establishes.
This guide translates four commonly blurred terms into a practical review: facility registration, current good manufacturing practice (cGMP), product claims, and FDA approval. It then shows how to read a Certificate of Analysis (CoA) as one document in a broader quality file and how to audit the language on a product page.
Four terms, four different meanings
Facility registration records a facility
Facilities that manufacture, process, pack, or hold food, including dietary supplements, generally must register with FDA unless an exemption applies. Registration gives the agency information about covered facilities; it is not an FDA endorsement or an approval of the facility’s products. FDA also does not issue a food-facility registration certificate as proof of product approval. (Registration of Food Facilities and Other Submissions)
If a brand uses the phrase “made in an FDA-registered facility,” it should be able to identify the actual facility and substantiate the statement. The sentence should not be paired with an FDA logo, a fabricated certificate, or visual treatment that suggests the agency evaluated the product. A registration record answers a limited administrative question: whether a facility is registered in the relevant system. It does not establish that a particular supplement is safe, effective, or compliant in every respect.
cGMP is a set of operating requirements
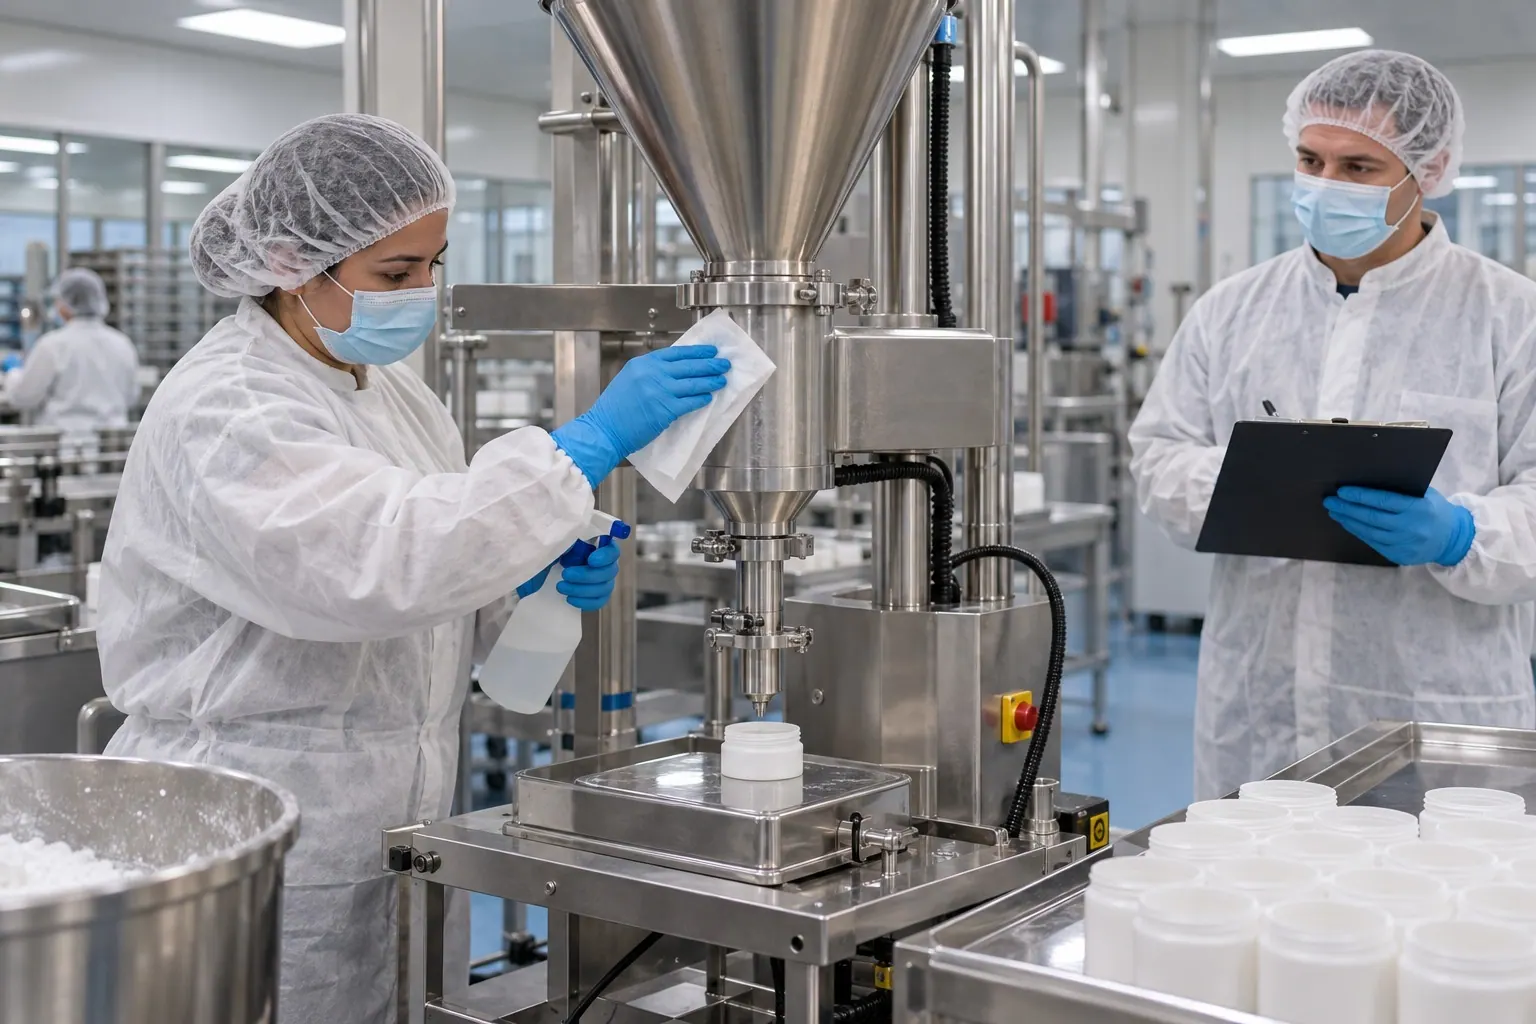
Under 21 CFR Part 111, covered people and operations involved in manufacturing, packaging, labeling, or holding dietary supplements must follow applicable current good manufacturing practice requirements. The rules cover quality-control responsibilities, specifications, production records, testing, and handling across the manufacturing process. FDA inspects facilities periodically; an inspection is not a permanent certificate that every product or lot meets every requirement. (21 CFR Part 111)
A third-party audit or certification may offer useful evidence, depending on who performed it, what standard and scope it covered, and when it occurred. It is not an FDA approval. “GMP certified” is therefore incomplete as a trust claim unless the brand can name the certifier, standard, scope, and date, and explain what the certification does and does not cover.
Private-label arrangements add a practical diligence step. Part 111 applies to the operations and responsible parties described in the regulation; contracts can allocate work between a brand, platform, and manufacturer, but a contract does not erase duties that the law assigns to a covered party. Separately, FDA says firms that manufacture or distribute supplements are responsible for ensuring their products comply with applicable law. A founder should map who makes, labels, holds, imports, and markets the product, then obtain records from each party for the responsibilities it actually performs. Do not assume that the founder automatically carries every manufacturing duty or that outsourcing removes every obligation.
Claims depend on the words and context
A supplement’s label and advertising must not be false or misleading. A structure/function claim describes an effect on normal body structure or function—for example, the role of a nutrient in maintaining normal bone structure. Under U.S. rules, the responsible firm needs substantiation, the required disclaimer must appear on the label, and FDA must be notified no later than 30 days after first marketing the product with the claim. A disclaimer does not rescue a disease-treatment claim or unsupported wording. (Notifications for Structure/Function and Related Claims in Dietary Supplement Labeling; FDA Issues Letter to Industry on the DSHEA Disclaimer; Health Products Compliance Guidance)
A claim that a product diagnoses, treats, cures, or prevents a disease can cause the product to be regulated as a drug. Context matters: the product name, images, testimonials, surrounding copy, and the overall presentation can all affect the message a reasonable reader takes away. Founders should review the whole page and its linked promotional material, not only the sentence printed beside the Supplement Facts panel.
Some health claims require FDA authorization before use. That is a specific claim pathway; it does not mean that FDA has approved the supplement carrying the claim. Keep the exact claim, its intended audience, supporting evidence, disclaimer, notification record, and any required authorization together in a claims file.
“FDA approved” is not a supplement quality shortcut
FDA does not approve dietary supplements or their labels for safety and effectiveness before they are sold to the public. Certain requirements may apply before marketing in particular circumstances—for example, notification for a qualifying new dietary ingredient or authorization for a specific health claim—but those steps are not blanket product approval. (FDA 101: Dietary Supplements; Questions and Answers on Dietary Supplements)
So if someone asks whether a dietary supplement is “FDA approved,” the accurate response is that supplements do not go through the drug-style premarket approval process. Avoid claiming or implying that FDA approved the product, its label, or the facility that made it. The distinction is precise and still clear to a customer.
What a Certificate of Analysis can—and cannot—show
A CoA can report test results for a named sample or lot. Its value depends on details: the product and lot identity, the sample represented, tests performed, methods used, results and units, acceptance specifications, laboratory identity, and who reviewed or released the lot. Ask whether the laboratory is independent and appropriately qualified for those methods; “third-party tested” alone does not answer that question.
Compare the result with the specification rather than treating a number as self-explanatory. Check whether the test measures ingredient identity, potency, or a relevant contaminant; whether its detection limits are suitable; and whether the CoA covers the finished product or only an ingredient. A report for one lot does not automatically speak for later lots. A certificate with no matching lot number, unclear method, or no defined specification leaves important gaps.
A CoA is one record, not the entire quality story. It does not by itself prove clinical effectiveness, validate advertising claims, demonstrate that every batch is equivalent, establish regulatory compliance, or guarantee safety for every consumer. Depending on the product and the question, a founder may also need specifications, batch and release records, supplier qualification, deviation and complaint handling, stability information, and applicable adverse-event procedures. The exact records depend on the parties’ roles, the product, and the regulatory requirements that apply.
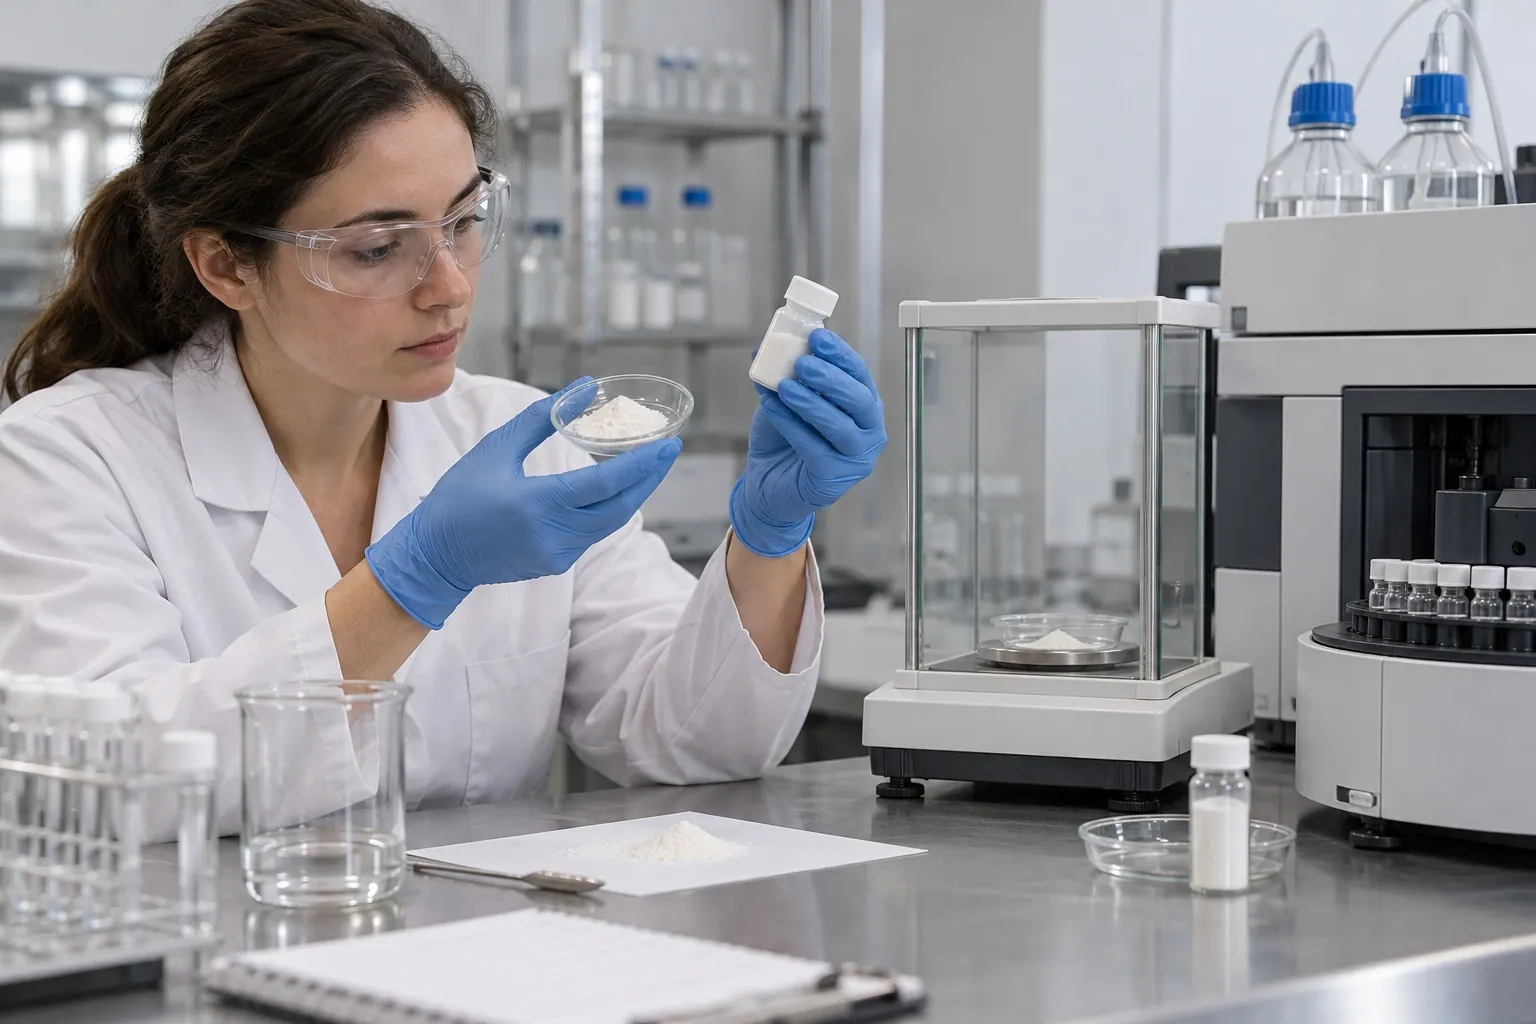
Before making a public statement based on a CoA, ask: Is this the finished product? Which lot does it cover? What did the test measure? Who performed and reviewed it? What specification defines an acceptable result? If the answer is unclear, narrow the marketing language or obtain the missing documentation.
A product-page audit founders can repeat
Start by copying every sentence about FDA, registration, certification, manufacturing quality, safety, benefits, or results from the label, product page, ads, creator scripts, and testimonials. Review visuals and context too: a badge or seal can imply endorsement even when nearby copy is technically qualified.
For each statement, record four things: who is responsible for it, what exact claim a reader could understand, which evidence supports it, and which rule or procedure applies. Then use this short review:
- Registration: Can the brand substantiate the facility and the wording? Does the presentation avoid implying FDA endorsement?
- Manufacturing: Which entity performs each covered operation? What Part 111 records or other evidence apply to that role? If a certification is cited, what is its named scope and date?
- Claims: Is the wording a structure/function claim, a disease claim, a specific authorized health claim, or another type? Is the support appropriate for the exact claim and audience? Are required label statements and notifications in place?
- Testing: Does the CoA match the product and lot? Are the methods, specifications, results, and laboratory clear? Does the public statement stay within what was tested?
- Approval language: Does any wording, badge, testimonial, or image suggest FDA reviewed or approved the product? Remove or correct that implication.
For example, “made in an FDA-registered facility” requires evidence of the facility’s status and must not be framed as product approval. “Manufactured under cGMP requirements” should be supported by the relevant operator’s records and role-specific evidence. “FDA approved” should be removed when used to describe a dietary supplement. “Supports bone health” needs review as a structure/function claim, including substantiation, the applicable label disclaimer, and timely notification.
Precision builds trust when the evidence is visible
Compliance language is most credible when it is specific and proportionate to the records behind it. A founder who can explain the difference between registration, manufacturing controls, test results, and approval can answer customers without turning an administrative entry into a quality promise.
The operational payoff is a cleaner product page and a better conversation with a manufacturer: ask for the record that answers the question, tie it to the responsible party and product lot, and make only the statement that the evidence supports. That discipline makes the brand easier to review and helps readers distinguish a documented process from an impressive-sounding badge.